Service Area
Saudi Arabia FDA (SFDA) Authorized Representative (AR)
Medical Device manufacturers without a local entity in Saudi Arabia must appoint an Authorized Representative (AR) to manage regulatory compliance.
SFDA Rep
When is a Saudi Arabia Representative Required?
For foreign manufacturers without a physical legal entity in Saudi Arabia, it is mandatory to appoint a local legal entity as their Authorized Representative (AR) to register and sell products. This is mandatory to comply with Saudi Food & Drug Authority (SFDA) regulations.
There must also be a formal, documented agreement between the manufacturer and the AR, approved according to Saudi regulations. This agreement should specify:
- The activities the AR will perform on behalf of the manufacturer.
- The types or groups of medical devices covered.
- Post-market surveillance obligations the AR must fulfill.
- The duration and termination terms of the agreement.
While manufacturers can appoint distributors as ARs, it is often recommended to have an independent AR to avoid conflicts of interest and maintain control over product registrations and regulatory obligations.
Responsibilities
Responsibilities of the Saudi Arabia Rep:
Liaison with the SFDA
Your AR serves as your official point of contact with the SFDA. All regulatory communications are handled through the AR. This includes responding to authority questions, coordinating audits or inspections, and ensuring that any regulatory updates or requests are addressed promptly.
Registration
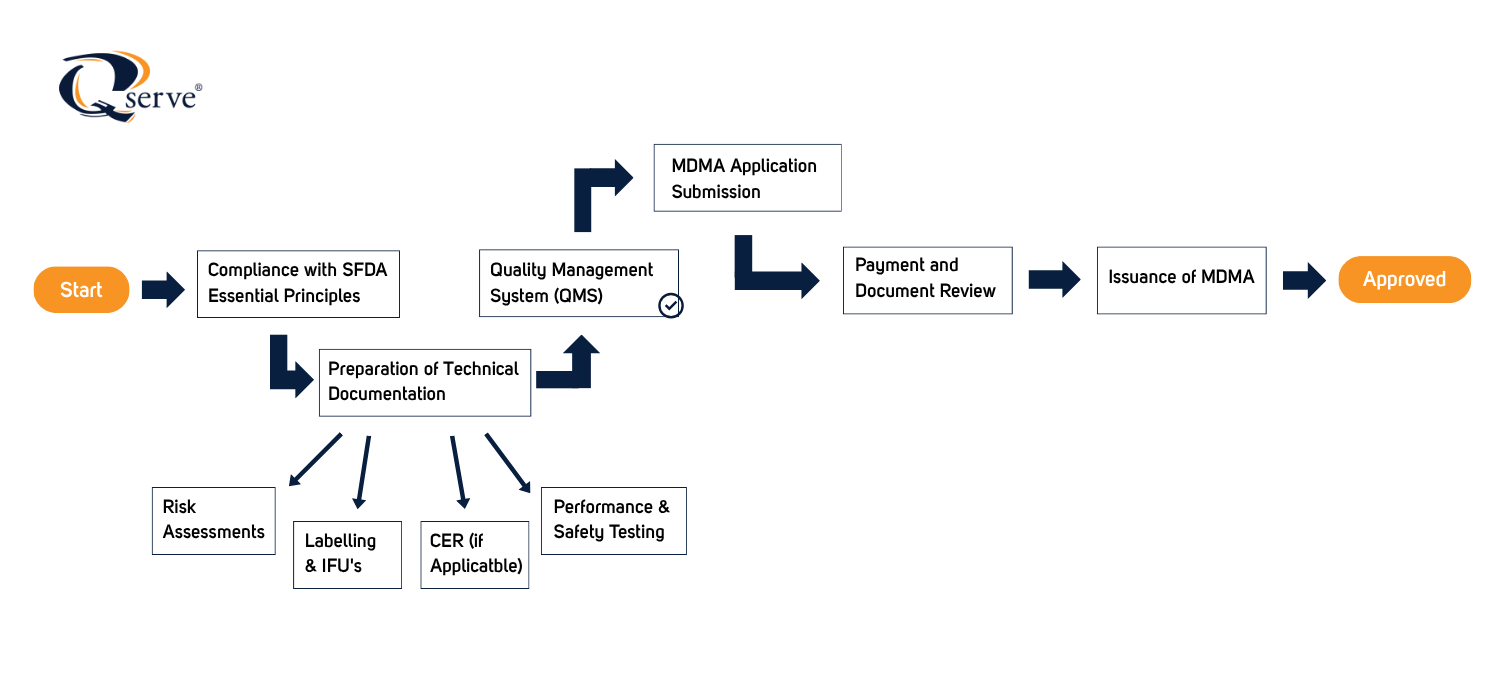
The AR is responsible for managing and submitting all medical device and IVD registration applications to the SFDA, including the coordination of the full registration lifecycle. The AR also tracks application progress and manages authority feedback.
Documentation
Your AR is responsible for preparing, reviewing, and submitting all required regulatory documentation in accordance with SFDA guidelines. This includes compiling submission dossiers, maintaining technical documentation and ensuring documentation aligns with Saudi regulatory requirements.
Compliance
Beyond initial approval, the AR ensures that all devices placed on the Saudi market remain compliant with applicable SFDA regulations and standards. This includes verifying labelling requirements and ensuring adherence to Saudi-specific regulatory frameworks. Labelling must be in Arabic and/or English with specific content requirements. The AR also oversees post-market compliance activities, such as vigilance reporting, complaint handling, and corrective actions when necessary.
MDMA submission
The AR is responsible for preparing and submitting the Medical device Marketing Authorization (MDMA) application. This process requires detailed technical documentation, evidence of safety and performance, and proof of compliance with applicable standards. The AR ensures that the MDMA dossier is complete, accurate, and aligned with SFDA expectations.
Currently looking for a service provider to support registration and Authorized Representative services in Saudi Arabia?
Blogs
Latest blogs
.png)
Know if your device needs a clinical investigation under MDR, and how ISO 14155:2026 shapes the study. Learn the exemptions, pitfalls and next steps.
.png)
Missing the 28 November EUDAMED deadline puts your devices at risk of non-compliance. Learn what’s at stake and how to get your data submitted in time.
.png)
Learn how to structure a PMCF plan under the EU MDR. Walk through sections A to G of MDCG 2020-7 and avoid four common pitfalls before your Notified Body does.